LX2006 LX3010 PRECLINICAL
CANDIDATES
Discovery
Candidates

Friedreich
Ataxia

Friedreich ataxia (FA) is a rare, hereditary disease caused by a pathogenic mutation in the FXN gene, leading to mitochondrial dysfunction. FA affects ~5,000 individuals in the United States and ~15,000 individuals worldwide. While historically recognized as a neuromuscular disorder, FA is a complex and devastating multisystem disease, with cardiac complications being the most common cause of premature death.

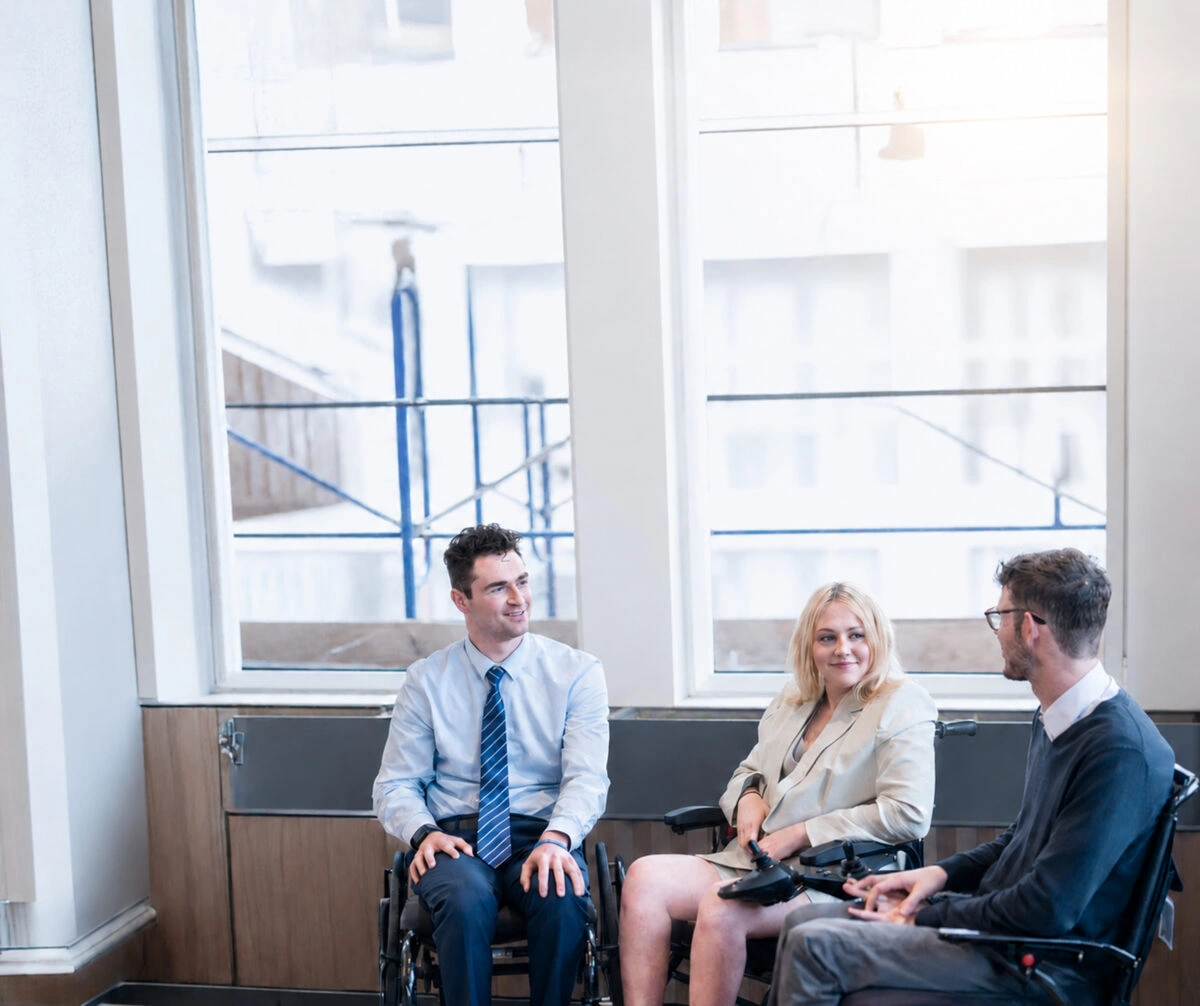
Liam, Hayden, and Christian, individuals living with FA
Programs: Indication: Target Gene:
Pre-clinical: Discovery Pre-clinical
Clinical: Phase I/II Phase II/III
Friedreich Ataxia programs
Pre-clinical: Discovery Preclinical
Clinical: Phase I/II Phase II/III
LX2006 Friedreich ataxia cardiomyopathy Gene therapy FXN
Pre-clinical: Discovery Preclinical
Clinical: Phase I/II Phase II/III
LX3010 Friedreich ataxia Small molecule FXN
Pre-clinical: Discovery Preclinical
Clinical: Phase I/II Phase II/III
PRECLINICAL
CANDIDATES
Friedreich ataxia Multiple modalities FXN
Pre-clinical: Discovery Preclinical
Clinical: Phase I/II Phase II/III
DISCOVERY CANDIDATES Friedreich ataxia Multiple modalities FXN

Disease
Overview

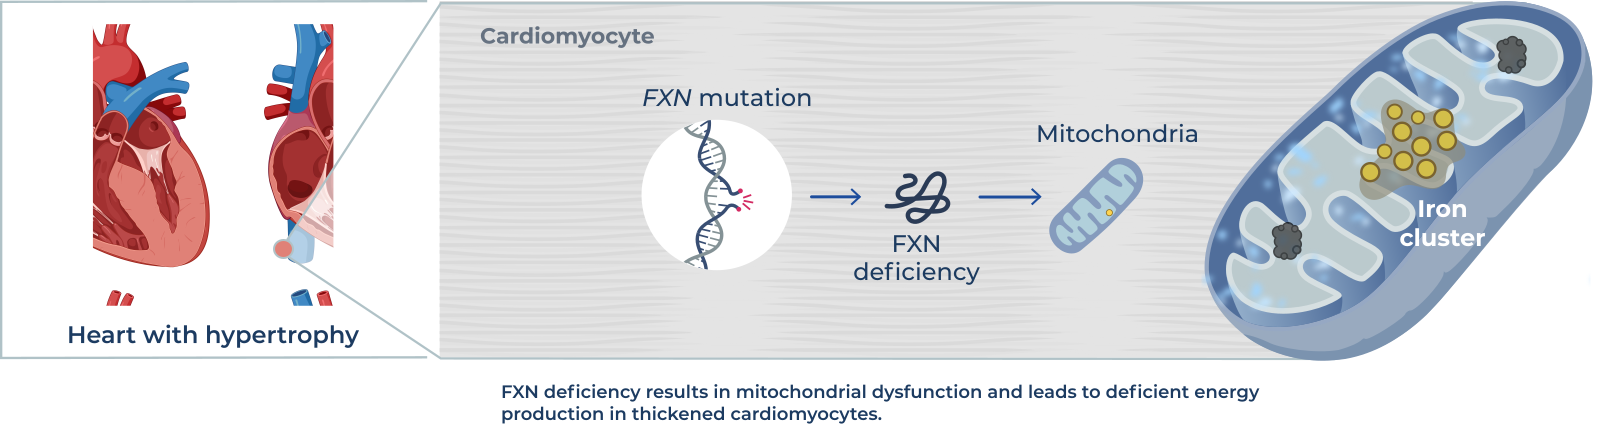
Friedreich ataxia is a rare, hereditary disease due to a pathogenic mutation in the FXN gene that disrupts the normal production of the frataxin (FXN) protein.

Friedreich ataxia is inherited in an autosomal recessive manner, where both inherited genes are abnormal, and symptoms usually begin in childhood. Absence of fully functional frataxin leads to damage to peripheral nerves and the parts of the brain that control movement and balance, leading to neurological symptoms that worsen over time, including ataxia or impaired muscle coordination.

Frataxin is also critical to the function of mitochondria and to the maintenance of cardiac function. As FA progresses, patients typically experience various heart conditions, including thickening of the heart muscle, arrhythmias, elevated biomarkers, progressive ventricular remodeling, and subsequent heart failure. Cardiac complications account for up to 80% of deaths in FA.

There is no approved treatment for FA cardiomyopathy (FA-CM), creating significant unmet need for individuals with FA-CM.

LX2006
Construct

LX2006
mechanism

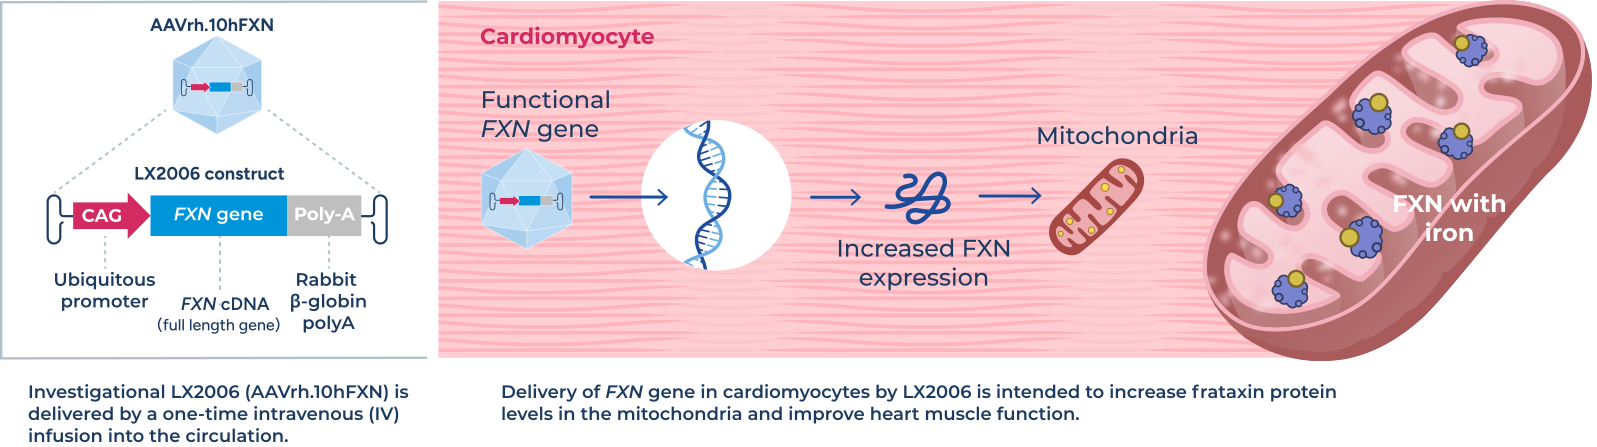
We are developing LX2006 as an AAV-based gene therapy delivered intravenously for the treatment of FA cardiomyopathy.

LX2006 is designed to transfer the FXN gene to myocardial cells and increase frataxin levels in the mitochondria. The increase in frataxin levels in the mitochondria restores mitochondrial function and energy production in cardiac myocytes.

LX2006 clinical data show sustained or deepening improvements across both cardiac and neurologic measures of FA.

LX2006 Regulatory Designations and Programs

Breakthrough Therapy Designation (FDA)

Rare Pediatric Disease Designation (FDA)

Regenerative Medicine Advanced Therapy (RMAT) Designation (FDA)

Chemistry, Manufacturing, and Controls Development and Readiness Pilot (CDRP) Participant (FDA)

Orphan Drug Designation (FDA)

Orphan Medicinal Product Designation (EC)

Additional Pipeline Candidates for FA

Lexeo is also evaluating several clinical, preclinical, and discovery candidates for FA.

LX3010 is a clinical stage, oral combination therapy designed to increase frataxin expression while addressing mitochondrial function and oxidative stress through complementary mechanisms (HDAC inhibition and NRf2). Early clinical data includes an approximately 6-point improvement in mFARS scores at 16 weeks and a mean nine-fold increase in frataxin protein levels from baseline in muscle biopsies across 11 FA patients.

Lexeo is currently evaluating two preclinical programs for the treatment of Friedreich ataxia (FA). LX3030 is an oral, tissue-penetrant small molecule (sub-500Da) designed to increase production of endogenous frataxin in the central nervous system. This third-generation benzamide HDAC inhibitor builds on published clinical work demonstrating oral benzamide HDACs increase frataxin in FA patients by epigenetic modulation and acetylation of chromatin. Preclinical studies of LX3030 have demonstrated robust increases in frataxin, supporting continued evaluation of LX3030 as a differentiated oral therapy.

LX3050 is a recombinant human frataxin fused to a proprietary anti-TfR1 Fab, intended to directly replace the deficient frataxin protein and utilize a TfR1-targeting brain shuttle designed to cross the blood-brain barrier and increase delivery of frataxin to the brain. In vitro studies have demonstrated dose-responsive improvements in measures of mitochondrial function, providing early support for the program’s proposed mechanism.

Lexeo is evaluating additional therapeutic modalities for Friedreich ataxia (FA) to treat the multisystem manifestations of the disease. Research is currently underway to evaluate a discovery stage ASO Fab conjugate program designed to stabilize FXN mRNA and thereby increase translation of endogenous frataxin protein. Lexeo is also evaluating cerebellar-targeted sequential dosing of frataxin gene therapy, designed to be complementary to systemically administered LX2006. This research is evaluating both intra-cisternal administration as well as a novel, intravenously administered, blood-brain barrier crossing capsid that is expected to enable a less invasive route of administration to the central nervous system following initial systemic administration of LX2006.

Lexeo will continue to evaluate these programs and will prioritize the opportunities demonstrating the strongest potential for clinical patient impact and regulatory success.
Continue the story
Learn more about arrhythmogenic cardiomyopathy and LX2020
EXPLORE ACM